Lyophilized peptide packaging changes: vial, stopper and seal review
Review lyophilized peptide vial, stopper and seal changes using a comparability plan for packaging identity, compatibility and documented change control.
5 min read · Sources checked 2026-10-22
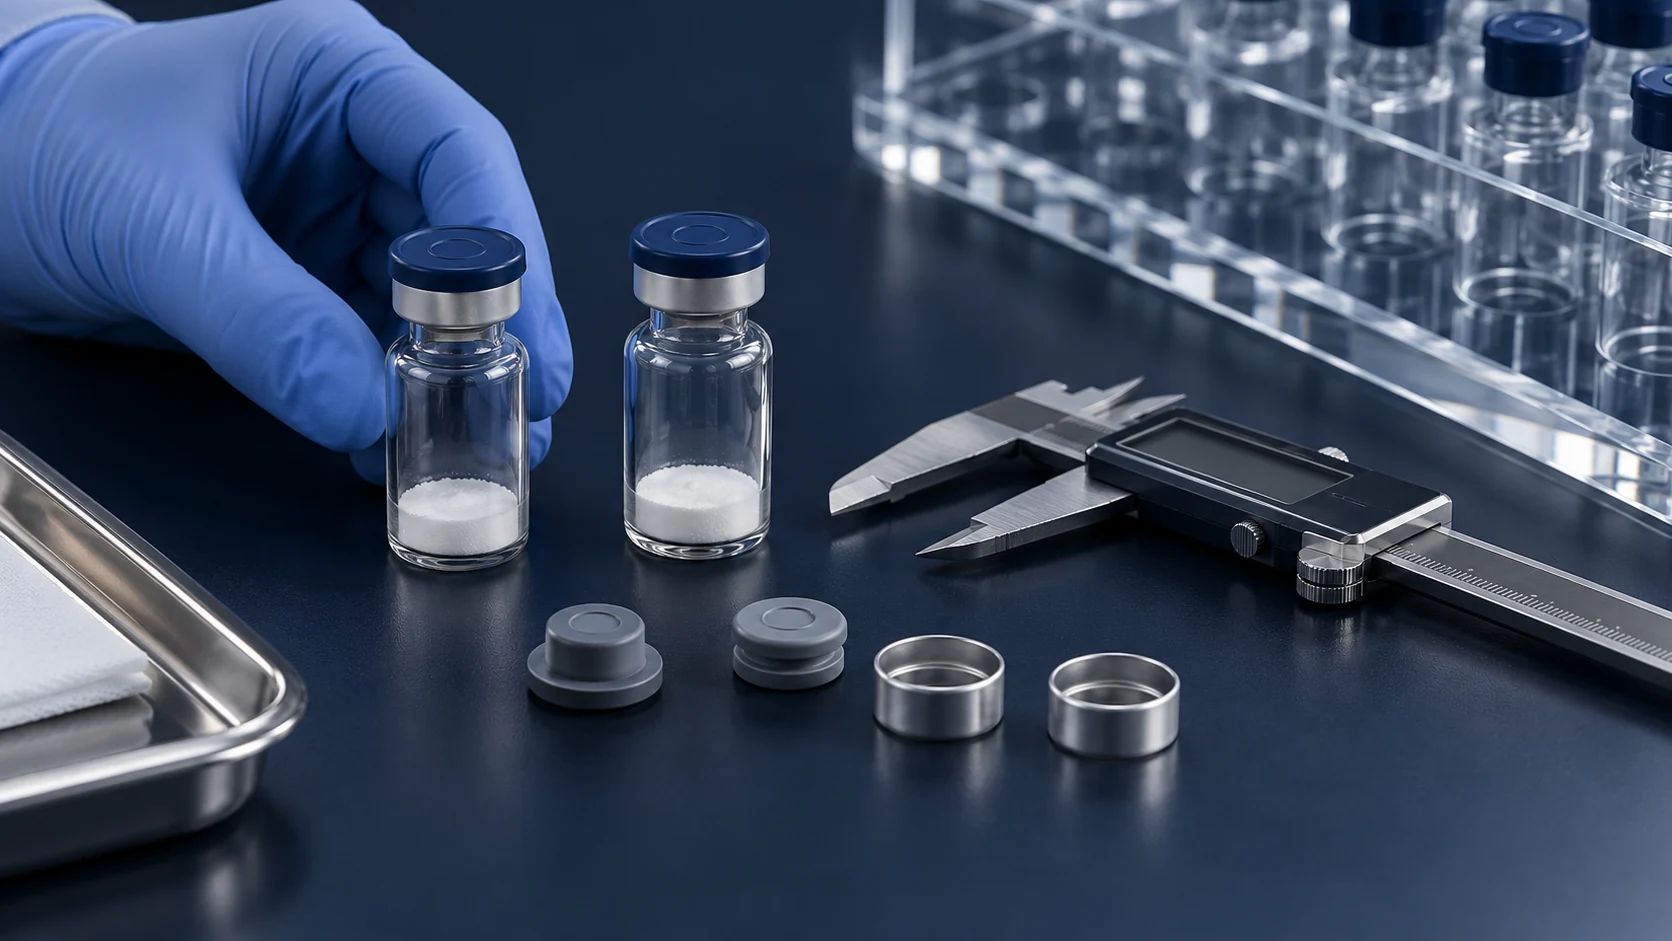
A change to the vial, stopper or seal should trigger a review of the assembled package. Identify the changed component, the material attributes it could affect and the evidence needed to accept the change. A matching external appearance cannot settle those questions.
Describe the complete container-closure system
Purchasing records often reduce a package to its nominal vial size and cap colour. For a lyophilized peptide project, the material-contacting surfaces, closure fit and assembly history also belong in the specification. A nominal 2 mL vial description states a container capacity; it does not establish compatibility with a particular stopper.
FDA’s container-closure guidance organises packaging suitability around protection, compatibility, safety and performance. [1] The guidance concerns human drugs and biologics. Here, those headings provide a way to organise questions for a research-material project; they do not confer a regulated product status on a research vial.
Separate product-contact parts from presentation choices
The vial and stopper can contact the material or its internal environment. A crimp holds the closure assembly in place. A flip-off top, label and outer carton have their own functions. An artwork revision may therefore require a different review from a new elastomer formulation, even when both changes arrive in one packaging order.
Record component manufacturers and part references where they matter to the agreement. A purchasing description such as “same blue cap” is too broad to establish that the glass, stopper and assembly remain equivalent.
Map the change to a possible material or package effect
| Proposed change | Question for the technical review | Record to request |
|---|---|---|
| Glass manufacturer, composition or geometry | Could the contact surface or closure fit change? | Component identity, drawing and applicable comparison evidence. |
| Stopper formulation, coating or dimensions | Could compatibility, contact behaviour or sealing performance change? | Material description, dimensions and a justified compatibility assessment. |
| Crimp component or assembly operation | Does the assembled closure still perform as specified? | Assembly specification and relevant package-performance evidence. |
| Label or carton construction | Could handling, identification or protection in distribution change? | Approved artwork, label checks and packaging review. |
The same review should state what remains unchanged. Keep the peptide form, presentation and comparison basis visible so that reviewers can separate a packaging effect from a simultaneous material change.
Distinguish extractables, leachables and physical compatibility
A material-contact assessment should address the interaction that matters for the actual product and package. Questions can include substances associated with packaging components, interactions with the material and changes in the closure’s physical behaviour. The FDA guidance discusses component composition and its influence on compatibility and protection. [1]
For procurement, ask the technical team to explain why the selected evidence fits the project. A generic component datasheet may help identify the material, but a supplier should explain how that information supports the intended use. The amount of evidence should follow the change and the agreed application.
Define what package integrity means for the project
Ask which package property the comparison will assess and how the result will be interpreted. An attractive crimp or an intact-looking cap is a visual observation. Any claim about closure integrity requires evidence appropriate to that claim. A research-use label also supplies no evidence of sterile manufacture.
Agree a comparability plan and its decision owner
ICH Q10 describes a pharmaceutical quality-system model that includes managing changes. [2] A research supply agreement can apply the same documentation discipline: describe the proposal, assess the possible effects, agree the review and keep the approval with the relevant specification.
- Identify the old and proposed component versions and explain the reason for the change.
- Name the material attributes and package properties that could be affected.
- Agree the comparison evidence, its acceptance criteria and the responsible reviewers.
- Assess whether existing storage or stability conclusions still apply to the revised package.
- Record approval, implementation date and the first affected material lots.
Where information is missing, record the open question and the owner who will resolve it. A signed change record should explain the conclusion well enough for a repeat-order reviewer to understand it without reconstructing the original email thread.
Put change notification into the supply brief
Request notification for changes that affect the agreed component identity, material-contact surfaces or package specification. State which changes need technical approval before implementation. Keep a copy of the approved package description with each applicable quotation and lot record.
For a new presentation, use the lyophilized powder OEM brief to define the package early. Review stability evidence and shelf-life questions separately, and connect accepted changes to the batch documentation set.
Frequently asked questions
Does using the same vial volume make two packages equivalent?
No. Nominal capacity does not establish the same glass composition, dimensions, stopper fit or assembled-package performance. Define the parts and the comparison evidence in the package specification.
Should a cap-colour change require the same review as a stopper change?
The review should follow the function and possible effects of the change. Cosmetic presentation changes and changes to a material-contacting elastomer may require different evidence.
Can an old stability statement be reused after a packaging change?
The responsible reviewers should assess whether the existing evidence still applies. Document the changed component, the possible effect and the justification for using or supplementing the earlier evidence.
Which lots should a packaging-change record identify?
Record the approved component and specification versions, implementation date and first affected lots. Keep enough information to connect a later enquiry with the package actually supplied.
Sources and further reading
- FDA: Container Closure Systems for Packaging Human Drugs and Biologics
- FDA/ICH: ICH Q10, Pharmaceutical Quality System
Sources checked on 22 October 2026. External references support the stated definitions and examples; they do not certify our products or facilities.